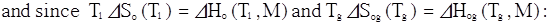jeudi 12 août 2021

.......................................... IT'S ABOUT TIME


                                                       IT'S ABOUT TIME! 


…about time for what?


No, no, no…this post is really about TIME, you know, the concept of time, the tic-tac of the watch, the flow that carries us along in our existence.

In physics, we measure time with respect to a reference, the second, and we express the flow of time by a multiple of that reference.  For instance the Big-Bang occurred 4,229 trillion seconds ago (13.4 billion years), and the life span of a human being, if everything goes right, may reach 3.16 billion seconds (100 years), enough to be part of a lot of good things and bad things…

So, time is usually perceived as an index of the flow of something passing by that we can measure. The consciousness of the reality of the flow of time is due to our perception of the differences between the past, the present and the future. The events which occurred in the past appear frozen like the objects in a picture or in a recorded movie that can only play the same sequences of the events, inalterably. There is a great certitude that it happened, one way and no other way, even if it’s difficult to reconstruct which way it was.

The present is the living reality of time: it is more difficult to apprehend because it requires defining a scale to gauge the dynamic changes occurring simultaneously and the magnitude of their interactions: what is the present for a rock, for a vibrating cesium atom, or for a living cell? The present is our reality but it is multiple: it could be light years, days or an hour, a second, a picoseconds (10-15 s) or even the Planck’s time, 10-43 seconds, the theoretical smallest time duration between the beginning and the end of any subatomic event.

The future is imaginary, or at least imaginable to physicists and engineers, because bound to the present, starting at the end of the present, generated by it (the famous initial conditions of a physical system), and perhaps “merging with the present”(uncertainty principle) at the Planck’s scale.

Time appears to be linked to the description of changes occurring to events, such as in the diffusion equation of heat exchange or in the propagation of sound or light in space. This is a passive role, the index role of a stop watch.

No changes, no need for the concept of time, no difference between past, present and future. Understood!  But how do we measure “no changes”? The measurement of something implies a duration, thus a change of time, an apparent catch 22!. Yet we know how to solve this problem: by the use of the concept of derivative in infinitesimal calculus.  The variation of the changes occurring to a variable, dv, is found by extrapolation to zero of an increment of time, dt:

                v’ = limit of dv/dt when dt→0 .

Each reality change is defined by v’, the ratio of 0 to 0 as dt becomes 0. This is an extraordinary weird thought that our description of reality, of our present, is made up of a series of 0/0! But the magic is that this works: we can describe natural events at no changes, neither time nor anything else: this is a timeless solution, time is just an index in such a mathematical construction.  

The question, though, of the physical legitimacy of extrapolating to 0 remains vivid: are we introducing a fundamental error of comprehension of physical phenomena by extrapolating to 0? Is this why quantum reality, described by the physics of derivatives, has such a hard time to make sense? I like to call the analysis surrounding this question: the structures of zero.  It involves describing the vacuum, of course, both the vacuum separating clusters of matter in the galaxies and other skies and the vacuum inside the atoms, inside all subatomic particles. Vacuum versus vacuum: the structures of vacuum.

Many important equations of physics are expressed as direct or partial differential equations to calculate the changes occurring to a parameter as a function of time. Time reality has disappeared in such expressions because of the use of the derivatives (1st order, 2nd order etc.): Time is totally passive in these expressions.   In other equations, such as in Kepler’s 2nd law of planet orbital motion, time is directly embedded in the description of the event, not its derivative: it is proportional to the surface area swept by the radius between the planet and the sun localized at the focus of the ellipse. Time is still a passive variable in all these equations, it is there to measure changes occurring to active variables.

In 1905, Einstein’s relativity made time and space correlate dynamically as if time was an active variable, not passive.  Einstein made clear that time, space and mass (of matter filling space) could not be considered independently of each other when motion was observed.  Einstein led the way to a forward vision, making time stretchable or shrinkable as matter moved at speeds near the speed of light: this was, indeed, a revolution in thinking!

However, is time a true active variable in Einstein’s physics when time (multiplied by the velocity of light) is simply used to create a four dimensional  timespace where the unit referencing time and the unit referencing space become linearly dependent?  In any case, Einstein’s new vision certainly bookmarked in our consciousness the end of a well established simple understanding of the concept of time.

What now? Could time be still misunderstood and play a REAL active role?  What does “an active role” mean?

If time comes as discrete Lagrangien durations, packets of “chronons”, say defined by a discrete set of Fourier frequencies and amplitudes, which we call  “the vertical structure” of this time duration, one can imagine that this structure of correlated chronons evolves as the event itself exhibits changes in time (for instance due to the acceleration of matter in timespace). As the vertical structure and the parameter describing the event are correlated, one can now talk about “an interaction” between the time structure of the chronon packet -representing the index to characterize the event’s evolution in time- and the parameter describing the event, say its electrical or magnetic properties, its position in space, its velocity or acceleration. This “interaction” makes time part of the event: the structure of the chronon packet and the total Lagrangien duration of the packet become an integral part of the event, interfering with it to optimize some aspect of the changes (minimize the dissipative energy). This type of interaction where the size of the unit and its structure are coupled with a minimum principle describes what I have called a “dissipative interaction”.  I have described such interactions in other disciplines of physics: dissipative interaction of Temperature and Voltage in the thermally activated relaxation of dipoles, and dissipative interaction between the conformational energy of conformers and their free volume to understand the visco-elastic behavior of polymeric melts in a novel way.

In other words, using the terminology above: is time dissipative? Is time visco-elastic?

If it is, all of our mathematical description of physics that now includes time as a passive variable in differential equations, for instance, should be revisited.

And if time is itself interactively part of what we measure, shall we continue to consider the velocity of light the ultimate reference to understand our past, by back extrapolation, not knowing the history of the structures of time that came along during the travel of the photons?

Our consciousness of time and space, realities elevated to the status of collective knowledge when they started to be defined and measured with respect to their reference unit (a concept generalized to all parameters considered in physics), is deeply engraved in all of us. This is ground zero of our collective elementary knowledge of physics. The measurement of these two fundamental realities, and of velocity as a corollary of their duality, marked a major step forward in our evolution as humans.

Yes, perhaps, IT’S ABOUT TIME to evolve from this 2,000 year old idea of a fixed reference unit to describe the fundamental parameters and constants of natural phenomena. Einstein led the way to such a forward vision, making time stretchable or shrinkable, but this was an “elastic” vision, also affecting mass and space elastically. We may have to think of time as a visco-elastic entity with an elastic component and a viscous (dissipative) component.

The direct impact of this new vision of time on our knowledge of the universe at the scale of our daily life might be insignificant, because time is an index under such conditions, and our mathematical  description of the laws of nature with such a passive time is clearly powerful and well adapted (infinitesimal calculus).  I propose, however, to go beyond a passive concept of time and articulate our comprehension of the interactions at the microscopic scale of quantum physics or at the scale of general relativity with help of a new mathematical description of an elasto-dissipative time in the equations.

Will the concepts of dark matter and dark energy survive such a dissipative description of the interactions? Will gravitation naturally rally the other fundamental forces?


Jean Pierre Ibar

jpibar@alum.mit.edu

University of the Basque Country


Arrosa, July 28 3021


dimanche 20 septembre 2020

WHY THE NEED OF A NEW RENAISSANCE IN (POLYMER) PHYSICS?


WHY THE NEED OF A NEW RENAISSANCE IN (POLYMER) PHYSICS?


Prof. Ilya Prigogine


The French word “renaissance” means “re-birth” in English, but, actually, it does not need to be translated: everyone uses the original French word to designate a new paradigm, a new era, a fresh start.

I used that word, Renaissance, yet added “New” in front of it.  Is it not a pleonasm to say: “a new renaissance”, like “a re-re-birth”?

Not if you consider that the 1st renaissance in polymer physics occurred with de Gennes, circa 1971-1979, introducing the reptation model of polymer dynamics.

The pre-de Gennes era, in polymer physics, was dominated by physical chemists: Bueche, Flory are typical examples, Treolar also pops in my mind, but there are so many others; I would need Boyer’s  legendary memory to be able to turn up a long list of names (see my previous blog #35).  

The pre-de Gennes era established the concept of macromolecules (Staudinger), and de Gennes School developed the scaling concepts to adapt the statistical mechanics of the small molecules to the dynamics of the long macromolecular chains. This field has become a very sophisticated mathematical model, indeed, after 40 years of fruitful advances due to ramifications and improvements.

Yet I maintain that we need to turn the page and start a new renaissance in polymer physics.  Do we need a clean slate?

Let’s roll back to the time when de Gennes started: how can we consider, differently, the statistics of interactions of the macromolecular chains?  After all, if the concept of macromolecule is not challenged, do we not need to select a single chain as our statistical system?  Like we always do in physics, once the properties of the system (here the single chain) is properly described, after accounting for the presence of the other chains that perturbs the property of a single isolated chain, we can extrapolate to the whole set of chains. This is the model that the reptation model proposed and elaborated for 40 years.

The clean slate that we actually need, in order to reformulate the interactions between the macromolecules, is not a small matter; in my opinion, it also bears general consequences in the way we should view interactions in physics, I mean statistically speaking.  The Boltzmann’s kinetic theory of gases inspired all current statistical models of the steady state of the interactions between a large set of units, molecules in the case of Boltzmann. This involves the description of a mean field calculated from the energy distribution function and it also involves a close statistics, where the canonical ensemble is well defined and constant.

Should the Boltzmann’s assumptions be put into question, challenged and perhaps even considered as THE problem to solve?  

If it is the case, not only do we need a new renaissance in polymer physics, but perhaps even, more generally, in physics.

You may not have recognized the man in the picture at the forefront of this blog. He is not as popular as Feynman, but perhaps should be. He is Professor Ilya Prigogine (1917-2003), Nobel Laureate in 1977 and author of several statistical books on the dynamics of “dissipative structures”[1], [2].

I happen to use almost the same words: elastic dissipative wave[3], dissipation energy, vertical structuring due to the minimization of the dissipative term, etc. in the description of the Grain-Field Statistical model of the interactions applied to polymers[4].  There is no doubt in my mind that the essence of my work on dissipation, started independently from Prigogine, has great resonance with what Prigogine has most brilliantly elaborated.  Perhaps luckily, though, I was not inspired by that work (at the time) and developed a different mathematical formulation of dissipation, possibly more adapted to the case of interactions between macromolecules, moreover mathematically simpler to apply, it seems.      

I am not going to elaborate these statements in a blog post, but since I have been working on those issues for the last 5 years and am ready to send to publishers a couple of books developing these new ideas on the subject[5],[6], I propose to “avant-premiere” a selection of general paragraphs from the books to illustrate what I have concluded and the questions that remain open.


A.  Excerpts from Prigogine1:


“We believe that we are only at the beginning of a new development of theoretical chemistry and physics in which thermodynamics concepts will play an ever increasing role.”

“In the classical theory of integrable systems, which has been so important in the formulation of quantum mechanics, all interactions can be eliminated by an appropriate canonical transformation.  Is this really the correct prototype of dynamic systems to consider, especially when situations involving elementary particles and their interactions are considered? Do we not have first to go to a non-canonical representation which permits us to disentangle reversible and irreversible processes on the microscopic level and then only to eliminate the reversible part to obtain well defined but still interacting units?”

“A general feature of interest is that dissipative structures are very sensitive to global features which characterize the environment of chemical systems, such as their size and form…, “

“For example, the occurrence of dissipative structures generally requires that the system’s size exceeds some critical value…”

“…It is precisely because of inequalities (3.2) and (3.4) that d2S is a Lyapounov function.  Its existence ensures the damping of all fluctuations. That is the reason why near equilibrium a macroscopic description for large systems is sufficient. Fluctuations can only play a subordinate role, appearing as corrections to the macroscopic laws which can be neglected for large systems.”


B   Excerpts from my books in footnotes 3, 5 and 6 (polymer interactions):


“…In our view, “conformers”, the constituents of the macromolecules, gather into statistical systems which go beyond belonging to individual macromolecules. A conformer is shown in Figure 7-1, duplicated from Ref. 276.  The macromolecules themselves represent a chain of "covalent conformers" put together as an entity.  The problem is to determine whether the chain properties, derived from its statistics, control entirely the dynamics of the collection of chains making up a polymer. This is what has been assumed by all the other theories, and this is what the Dual-Split kinetics and the Grain-Field statistics challenge…” 


“…to simplify, one could view the difference between our statistical model and the classical model to describe the properties of polymers as follows: according to the classical views, the statistical systems are the macromolecules, i.e. a network of chains; the properties of the chains are disturbed by the presence of other chains and by the external conditions (temperature, stress tensor, electrical field, etc.).  This classical definition of the statistical system contrasts with our approach where the statistical systems are the “dual-conformers”, not the macromolecules, assembled as a network of dual-conformers. The interactive coupling between the dual-conformers is defined by a new statistics, the Grain-Field Statistics, which explores the correlation between the local conformational property of the dual-conformers and their collective behavior as a dissipative network…”


“… the statistics that are used by the classical models and by our model to describe the RIS (rotational isomeric states) of the conformers are fundamentally different: the classical molecular dynamic statistics is the Boltzmann statistics, famous for its kinetic formulation of the properties of gases. The Dual-Split or Dual-Phase statistics, leading to the Grain-Field Statistics, is inspired by the classical Boltzmann concept but departs from it by defining a dissipative term in the equations and assuming that the Free Energy remains always equal to its minimum value, that of the equilibrium state, even for transient states. The kinetics created by such changes in the fundamental equations result in the formation of Free Energy structures, which we have once called “the Energetic Kinetic Dissipative Network of conformers (EKNET)” ([265] to [270])  and more recently, while dealing with rheology “ the Elastic Dissipative Network” ([276], [283a])…”


“…In our analytical formulation of the dynamics of these “open dissipative systems of interactions” generated by our two modifications of the classical formula, we realized that essentially two mechanisms of structuration of the Free Energy prevail and compete: a “vertical structuring” and a “horizontal” structuring, each specifically applying its own version of the basic equations. This distinction increases the complexity of the analytical solution but is, in our opinion, a fundamental aspect of the way interactions work. The vertical structuring refers to a split of the units (collectively interacting in the system) into 2 compensating sub-systems having each a different statistical partition. The horizontal structuring offers a different split of the collective set, via the generation of Ns identical sub-systems, each with the same statistical partition. Each split mechanism generates a dissipative function. The total dissipative function ought to be minimized (it is 0 at equilibrium), a condition that creates their compensation, i.e. whether they work independently, in sequence or together…”    


“…the details of the simulations performed using this model of polymer interactions shows that there is a temperature that we associate with TLL, that is the dynamic transition temperature beyond which the classical molecular models based on the Boltzmann statistics and our open dissipative network model are compatible and coherent. This stipulates that classical molecular theories of polymeric materials will provide the same results as our model for T > TLL, a temperature at which the dissipative function kinetically collapses.  Below TLL, the behavior that results from the interactions is dominated by the statistics of the Dual-Phase and Cross-Dual-Phases; thus, below TLL, the macromolecular aspect does not statistically dictate the properties. The projections of these macromolecular statistical models (reptation, for instance) are physically unfounded below TLL, in our opinion, which explains their failure to describe the experimental results under those conditions (Ch. 7 of [276], [283b])...”


“…If one tests the predictions of the classical approach under conditions that bring its state above TLL, one may conclude that those data validate the classical views since they provide correct answers in the range tested. This is not an easy task, because TLL is rate dependent, pressure and shear dependent and molecular weight dependent. Thus, although one will find in the literature convincing experimental evidence of success for the classical models, which is the reason for their acceptance, we claim that these successes are due to the use of conditions that bring the state of the polymer above its TLL transition…”


“… in most experimental set-ups used by the industry, TLL is raised to such high values, due to the high rates and pressures, that the range of validation of the classical macromolecular dynamics to predict the properties is in default: the use of such classical models in such conditions provides the wrong answers….”

“…we have advocated elsewhere (Ch. 7 of [276]), to abandon the classical interpretation of polymer physics by single chain macromolecular dynamics because of its inability to describe the full range of its behavior and other essential properties of polymers such as the dielectric TSD/TWD responses analyzed in [283f]…” 


“…we propose that the mechanism of relaxation, in polymers, is due to the dynamic coupling of two types of splitting processes of the total statistical population of conformers in interactions: the creation of Ns(t) Energetic Kinetic systems (horizontal splitting) and the modulation of the conformational structure of these systems by the dissipative function (vertical splitting)...”


“…It might be more appropriate to categorize TL,L as one of the kinetic manifestations resulting from the cooperative kinetic process already giving rise to Tb, ,Tg and Tg,ρ.  Beyond TL, L, the organization of the inter-intra molecular interactions between the various dipoles as a dissipative network is kinetically inefficient, hence has ended.  As we said earlier, a description of the properties of the polymer by invoking the properties of the individual macromolecules embedded in a mean field is acceptable from this point on…” 


C. Excerpts from the book in footnote 4 and in Vol. I of the book in footnote 6 (Grain-Field Statistics):


“…The study of kinetics is a discipline that describes the evolution of the units of a population of, say, chemical molecules that participate in chemical reactions. Another example would be to describe the evolution of units of a population which could occupy different “states”. Many other terms have been used to describe the same objective: “statistics”, or “dynamics”, for instance, as shown in the following definitions: the population partition that evolves with time can be studied with the tools of “statistics”, a transient statistics in fact, a field also regarded as “dynamics”.  All these definitions are used in our presentation. The important thing here is to define the terms quantitatively...”


“…Can we modify the set of equations driving the kinetics so the system Free Energy stays at its minimum value at all times?  The Dual Split Kinetics model describes new sets of kinetic equations fulfilling these conditions. There are two types of solutions that we have studied, vertical and horizontal splitting, and several possible hybrid combinations of the two…”


”…In this section we present the assumptions driving the new non-equilibrium statistics and study the difference between its results and results obtained classically. The new equations converge to traditional kinetic equations at long times or under "true" equilibrium conditions. Under non-isothermal conditions the system becomes self-dissipative, and the duality is responsible for a structure of the Free Energy…”


“…Note the presence of an additional term, Ln (Nb/Nf), in the expression of the Free Energy. This function is what we designate the "dissipative term"… “Its introduction is fundamental in our work on interactions; it is the source of the originality of the new statistics and results in the study of a new generation of dynamic open-self-dissipative systems…”


 “…In summary, simple relationships between Lnux, Δx and Δe exist which are revealed by varying Δe in Eqs. (5) to (7). The vertical splitting kinetics is, on its own, powerful enough to simulate the effect of activating the dipoles (permanent and/or induced) at the polarizing temperature Tp, and observing its thermally activated depolarization as a Debye current...”


“…We now imagine solutions that combine the Vertical and Horizontal Dual Split Kinetics to simulate the dynamics of statistical units in interactions… we just want to illustrate one of the solutions of the Grain-Field Statistics that we have explored extensively to simulate the thermal properties and the rheology of polymers. More generally, the description of the several combinations possible and their simulation constitutes a vast and fascinating program of investigation. Additionally, among the various solutions, the challenge is to recognize what combination could possibly correctly simulate the specific interactions in a given field of the physics of interactions, not just polymer physics…”


“…In each of the combinations of Vertical and Horizontal structuring mentioned above, we are dealing with auto-generated open dissipative systems driven by the energetic kinetics assumptions, i.e. by solutions of the structure of the Free energy  so that the minimization of the total Free energy, for the collective set, always remains equal to the equilibrium value at that temperature. An open system occurs when its total population is not kept constant. Such a system compensates its openness by modulating the structures of its Free energy at various levels (scales)...”


“…Figure B-21(below) provides another interesting perspective of the open systems dynamics when their total population, Bo, is submitted to a fluctuation around a mean value.  The system is again defined by Eq. (12), but we now suppose that the value of Bo varies with time like a sine wave as it is cooled  at rate q=-1 oK/s:

Bo= A+B sin (Ct) and T=To- qt.   A=1000, B= 100 are used in Fig. B-21. 

The problem is more complex to resolve numerically, but is still quite tractable. The solution for Ns (T) is plotted on the same graph as Bo (T) for comparison. As the temperature decreases, from right to left on the figure, (Bo(t) is represented by the squares, and Ns(t) by the open dots.

We see in Fig. B-21 that Ns varies periodically but nothing like a sine wave.

What is interesting, however, is to follow the period of the oscillation and the maximum and the minimum of the amplitude of Ns (T), from right to left: The amplitude of Ns has difficulty to rise up beyond its initial value at high temperature, showing a slow increase of the peak maximum value, and a strong non-linear oscillating systems, with lots of harmonics. . Beyond the 4th oscillation of Bo(t), however, the peak maximum value increases rapidly and levels off to a plateau value, and the oscillation becomes cleaner, indicating the muting of many harmonics. The other remarkable feature is the period doubling of the oscillation at lower temperature. At high temperature, the period of Ns(t) and Bo(t) are the same, but as we cross the transition temperature on cooling, the period of Ns(t) becomes twice the period of Bo(t), a phenomenon observed for “time crystals”, for instance.  

This Horizontal dynamic system is capable to produce a transition separating two temperature regions presenting very unique characteristics in material physics [283d]...”   


Figure B-21


“…This challenge of finding the correct combination of horizontal and vertical splitting to describe interactions requires a dedicated book [283d]… An important chapter of that book is titled: “The Dynamics of Open-Dissipative Systems of Interactions: the Question of their Finitude and Stability”.  This chapter, whose title, obviously, resonates with the work of Prigogine in the seventies [500a], although it totally differs from it for its mathematical treatment, should lay down the basic map of what needs to be resolved by the next generation of research scientists (interested in these solutions) to better understand how the local and the global interactions structure one another to generate transient and steady state events in time-space, i.e. describe what we call reality (Time vs Space, Matter vs Vacuum). This would offer new perspectives on the description of the different types of interactions, and of their unification as solutions of the network of open dissipative cooperative systems.  The 1st Renaissance in physics, from Newton to Einstein for gravitational interactions, to the Standard model of interactions to describe the infinitely small, may have exhausted its resources to complete a grand unification. Should the 2nd Renaissance, inspired by Prigogine [500b], now take the baton?...” 


Jean Pierre Ibar

New School Polymer Physics

jpibar@eknetcampus.com


September 14, 2020

Blog Post #36


[1] Prigogine I., “Time, Structure and Fluctuations’, Nobel Lecture (1977).

[2] Prigogine I., Nicolis G., “Self-Organization in Non-Equilibrium Systems”, Wiley (1977) ISBN 0-471-02401-5. Also:

Glansdorff P., Prigogine I.,”Thermodynamics of Structure, Stability and Fluctuations”, London, Wiley-Interscience (1971)

[3] Ibar J.P., “Physics of Polymer Interactions. A Novel Approach.  Application to Rheology and Processing”, Hanser, (2019). 

[4] Ibar J.P., “Grain Field Statistics Applied to Polymer Physics”, book in preparation.

[5] Ibar J.P., “Application of the Dual-Phase and Cross-Dual-Phase model of Polymer Interactions to the Rheology of Polymer Melts. Temperature and Molecular Weight Dependence: A New Approach.”, book, submitted for publication.

[6] Ibar J.P., “Interactive Coupling in the Amorphous State of Polymers”, books,  Vol. I and II, Accepted for publication 


jeudi 31 octobre 2019

A Tribute to R. F. BOYER (Post # 35)


STANDING ON THE SHOULDERS 
OF A VISIONARY GIANT:

RAYMOND F. BOYER


It took me more than 40 years to fully realize the extraordinary vision that Boyer had in believing and pushing forward, in his lectures and numerous publications, the importance of his TLL transition in the upper amorphous melt, and his Tg(U) and Tg(L) glass transitions in semi-crystalline materials. Both concepts were truly at odds with the established models of the amorphous state and of crystallization from the melt, at the time. Both concepts still remain, 50 years later, almost totally forgotten; in the grand library of the world knowledge, some would even rightfully say:
 “who needs that?”

This blog post #35 shows how to derive the existence of TLL, and how to explain it.  It also introduces the dual and cross-dual phase concepts that are behind the presence of the two Tgs in semi-crystalline polymers.

Thus, Dr. Boyer was right! TWICE!
            

This post is a brief description of my steps which led to the explanations of TLL and of the double Tgs.

Prof. R.F. Boyer inspired me to question my own knowledge in the field of polymer physics.

            The 1st time I met Prof. Boyer was at MIT in the spring of 1972 when he came to give us a lecture on his 2 Tgs in polyethylene: Tg(L) and Tg(U)[1].

            I already had heard of him from the classes I had attended at MIT, and of his controversial views regarding the existence of his TLL transition relaxation, located somewhere at ~1.2 Tg (oK) in the melt, which he had coined the “liquid-liquid” transition[2].

         After his conference, I had the privilege to be invited, in the office of my supervisor, to a one on one private conversation of an hour or so with him. Needless to say, this meeting had a profound impact on me. Not only was Ray Boyer a brilliant polymer physicist with an amazing memory capable of recalling and quoting all that had been published and considered important in polymer physical chemistry and physics in the previous 30 years, he was also engaging, kind and witty, humble with a  sparkling intelligence that made me instantly like him.

           The 2nd memory I have of Boyer took place in the summer of 1973 in Midland, Michigan, where he held the position of Director of Corporate Research at Dow. At a time when corporate research still blossomed from fundamental science, I am sure that he and Dr. Turner Alfrey were responsible for my being given the chance to live the experience of a summer job at Dow. Boyer continued to expose and inspire me with his new ideas and with his list of remaining problems to solve in polymer physics. 

          I almost exclusively focused on TLL, since my PhD research involved a new understanding of the interactions in the amorphous state[3]. I met Boyer’s wife and daughter, was invited to his place and even learned how to clear away the nasty smell off a dog sprayed by a skunk! I remember the poor dog in the bathtub being brushed with tomato juice all over its body!

            And then, I have a funny anecdote that came a few years after I had returned to France; this happened at a time when Ray Boyer had paid me a visit in my materials research lab in Ballainvilliers, near Orly airport, a place literally in the middle of nowhere, surrounded by huge farmer fields growing potatoes or who knows what else. We were invited for lunch at my home. 

          My car was a Renault 4L that I had bought for 500 Francs, i.e. less than 100 euros; a crusty white little thing that had the merit to drive through shortcuts on tractors’ tracks in the middle of muddy and slippery fields. Ray and I had started a passionate discussion on some aspect of the TLL transition, as I was driving through the fields, sliding here and there like on ice, not really paying attention to the driving. 

            At one point I realized that I was the only one talking, that Boyer was actually holding onto his seat with his two hands, his mouth open, intensely watching the wipers making noise and swiping mud traces on the windshield. It must have rained all night, or all morning, I also suddenly realized. “Are you OK”, I asked him. He looked at me and said, with a weird deep voice:

            “How far are we going? Are you sure we will make it?”.     

            These same questions seem to have framed my whole research life: How far am I going with my new ideas on interactions in physics? Am I sure I will make it?

         It was very promising yet surprising that a famous scientist such as Boyer showed such a genuine interest in my work. However, I was obsessed by the implications of the new ideas I was tossing in my head, and realized that my work had just started: that I needed to dig, to think, to learn…, yes, just learn and think…, intensely..,. and Boyer was a living library to learn from (BTW did you notice the size of his forehead in the Hall of Fame picture above…?).
 
   So, regarding my interest in Boyer’s controversial ideas, I was uncertain what to think of it: what was the impact of TLL and of the double Tgs on my own theoretical model of polymer interactions?  Should I discard these Boyer’s concepts…, like everybody else…, or did my model explain them?... Should my model explain them?... What if Boyer was right and my model did not explain them… (that was the case of the classical models…)?

      The truth of the matter is that, in my lab, when I kept repeating Boyer’s experiments and the analysis of his results (DSC, TSC, dynamic viscosity etc.), TLL was there, not very pronounced, but there, for sure. When I devised new ways to analyze the data (computerized double-shifting[4]), TLL appeared when it was not discernable before, or even became more prominent than before when it used to be a fuzzy signal.  I published these findings: no reaction; the TLL case was closed, Boyer had lost, Flory had won: no local order, no TLL, period!

         Then history turned the pages: de Gennes and his reptation ideas became the new paradigm to follow: forget Flory, forget TLL, move on, and, by the way, there was no place for a TLL transition in the melt from the stand point of polymer dynamics!  If reptation was right, no TLL, so move on!

        Still, in the back of my head, the real question remained: what was the meaning of TLL and what was its impact on the properties of the melt? Did it affect processing, for instance? Was TLL important? Was TLL the David who will make fall the reptation Goliath?

      I knew that the local order explanation offered by Boyer was probably incorrect[5], yet that Boyer could still be right that TLL was a real transition- relaxation.  I needed to understand it with my new theory on interactions. But how?


          Reader:  a brief fast forward to the present (it’s linked, you will see):

I authored a book, just published by HANSER (introduced at the K-2019 show in Düsseldorf):


The Physics of Polymer Interactions. A Novel Approach. Application to Rheology and Processing.


            You will encounter many references to the TLL transition in this book, each time showing a strong manifestation of its existence, in the middle of the melt temperature range above Tg, very near where Prof. Boyer had envisioned and predicted it (~ 1.2 Tg). This new evidence was obtained recently, using Rheo-Fluidizer processing equipment that shear melts under vibration to trigger various non-equilibrium states before they are frozen into pellets. The analysis of those treated pellets showed a strong TLL under recovery conditions. TLL was not only impossible to miss, but it could also be manipulated by the processing conditions, changing the vibration frequency or amplitude, or changing the pressure in the Rheo-Fluidizer.

           Yet, you will not see a mathematical demonstration of what TLL is, in that book. This QED and its explanation is given in Chapter 6 of another book which I have just finished, whose tentative title is:

Application of the Dual-Phase and Cross-Dual-Phase model of Polymer Interactions to the Understanding of Melt Rheology.

            Now, let’s return to our post story:
           
TLL is a real transition: QED.
 
Excerpts from Chapter 6 of that new book follow:

...
             After a thorough description of the interactions for M > Mc in the previous chapter, our objective, in this chapter, is to show that the dual-phase approach also provides a better understanding of the interactions for low molecular weight polymers (Mc
).

            In particular, the current theories of the melt liquid state have ignored, even vehemently opposed, the existence of a "Liquid-Liquid transition", TLL, above the Tg of polymer melts. The controversy has been introduced in the previous chapter in the case of the M> Mc data. We will now use a modified expression of the Vogel-Fulcher equation, which we call the "Thermo-Vogel-Fulcher equation”, to show that TLL naturally derives from it, demonstrating its fundamental aspect.  

            We also examine the rheological and thermal analysis consequences of the existence of TLL and cite the experimental evidence of TLL exposed by Boyer and others, ignored by the current theoretical models of the liquid state of melts, to show that this evidence may be interpreted and derived from the dual-phase and Cross-Dual Phase aspect of the interactions in polymers.

            Everybody knows the famous Vogel-Fulcher hyperbolic description of the temperature dependence of the Newtonian viscosity, log ho:
(1)


Now, let us define the Thermo-Vogel-Fulcher expression  (TVF) of the Vogel-Fulcher (VF) equation:
(2)
.


T1 is a “Reference temperature” of the TVF equation; such a reference state also appears in the WLF expression in its most general form.

The changing of the reference state in the TVF equation and its consequence on the value of DHo and DSo is similar to the variation of the WLF constants C1, C2 with the choice of the reference state, (T1, Logho1) in the WLF equation. The WLF constants are often quoted with a reference state taken at Tg. (Eq. 3):
(3)


This led Williams, Landel and Ferry to speculate that the C1g and C2g constants of the WLF equation were universal constants [2]. We have challenged such a claim in another publication [6], and it seems that most of the polymer scientists, our days, concur. When the reference is switched to T1, it is straightforward to find new WLF constants, in a way similar to what we will do in the following to find the evolution of DHo(T1,M) and DSo(T1,M) as T1 and M vary.

Our interest is to choose T1 such that the free volume remains constant for the reference state, as M varies, contrary to what is usually done when either T or M is alternatively changed as the other variable is constant, as if the effect of these two variables were independent. This is the main question raised and studied in this book: is it justified to separate the effect of M and T on the viscoelastic properties of polymeric melts?

Once the equations of DHo (T1, M) and DSo (T1, M) are known, it becomes straightforward to compare their values at constant free volume, i.e. at (Tg + A), where A is constant, and Tg is a function of M.

In the TVF equation, T∞ is also a function of M. In other words, we need to know both Tg(M) and T∞(M). This is given in Eq. (4):
 (4)


where ao and a1, are determined by Differential Scanning Calorimetry (DSC) at a constant heating rate, and bo and b1 by Vogel-Fulcher fits of the Newtonian Viscosity dependence on temperature. When T is expressed in oK, M in g/mole, MAJESTE (UPPA, PhD thesis, PAU-France) has stated that:
(5)

              ao= 371           a1=114,000
              bo=321.4         b1= 83,000

Derivation of the expressions of DHo(T1,M) and DSo(T1,M).

We start by rewriting the TVF equation at 2 reference temperatures T1 and Tg:
(6)


We now combine the 2 expressions of log ho(T):


We find ks=B. the numerator in the Vogel-Fulcher equation (1)

In summary:
(7)

    Note that T1 can be chosen arbitrarily in Eq. 7, whereas T∞ is determined by the value of M as given by Eq. 4. Some important values of T1 will be of particular interest in the following: T1=Tg, Tg∞, T1=Tg+A, and T1= T∞ +A’, where A and A’ are constant when M varies.
We will only deal with T1=Tg(M)+A in this blog post.


Study of the effect of M on DHo(T1,M) and DSo(T1,M) at constant Free Volume defined from Tg(M).

Instead of comparing the parameters of the TVF equation for  various M < Mc at constant T1, we  consider, in this section, comparing the parameters at constant free volume A, where A is established with respect to the Tg of the polymer factions, since, for M < Mc, Tg (M) is a strong function of M: T1= Tg(M)+ A.
From Eq. 7, we have:
(8)


When A is constant when M varies, the free volume, defined by (T1-Tg) remains constant .

We create a set of values of DHA(M) at A given, using Eq. 4 to determine Tg(M) and T∞(M).  We plot in Fig. 3 DHoA vs 1/M at constant A for various values of A varying between A=23 oC and 93 oC. We see in Fig. 3 that straight lines visually pass through the data, and the linear regression confirms that the linearity is rather good (last column of TABLE 2).  TABLE 2 gives the Intercept and the slope of the lines in Fig. 3, when A varies. What is interesting is that the slope starts from a positive value at T1=Tg (A=0), decreases as A increases, flattens out to 0 across all values of M for A approximately equal to 70 oC, and becomes negative as A continues to increase.  For reasons which will become apparent in the following, we will associate this peculiar temperature T1 that makes DHoA remain constant (to approximately 2600 in Fig. 3) across all M when the free volume is set at A~70 oC, as the TLL temperature of Boyer, the liquid-liquid transition.  This means that the state of the melt at that particular temperature is an iso-free-volume state with also a constant Thermo-Vogel-Fulcher Enthalpy, DHo: an iso-free volume-iso-enthalpic-sate, a unique situation as M and T1 vary.

                                    A          INTERCEPT      SLOPE              R2
                                           0          5213.46           14490364        A →0
                                    23        3806.02           677647.25       0.998
                                    46        3066.31           188407.88       0.9989
                                    70        2587.606         -39.6049          0.9768
                                    93        2284.95           -74180.24        0.9995

TABLE 2
Results of the regression lines in Fig. 3 of DHoA vs 1/M at A constant. A varies from 0 (Tg state) to 93 oC in this plot. PS data from Majeste [6].


Fig. 3
                                                                                                                                   
Finding the reason for the TLL transition.
PS Dynamic data of Majeste[6]-DHoA vs 1/ M at various A.

The TLL(M) is the temperature corresponding to the 0 slope, i.e. for A~ 70 oC.  TLL is an iso-free volume-iso enthalpic state. Graphically, we found :
TLL(M) =Tg(M)+70.5 oC  for monodispersed PS with M < Mc
Let us now find the mathematical expression of DHLL and ALL derived from the Thermo-Vogel-Fulcher expression of DHA(T1,M) at free volume constant T1=Tg+A
(9)


Numerical Application (Majeste’s data PS monodispersed M < Mc)
(10)
            B=703.5571; ao=371 oK, a1=114,000; bo=321.4 oK, b1=83,000

Therefore:
(11)

            ALL= 70.441 and DHLL= 2587.275

confirming the values found empirically in Fig. 3.

In other words, when T1=Tg+ALL, regardless of the value of M, DHLL=2587.275=constant.

The TLL is an iso-free volume and iso-enthalpic state of the melt, a very unique situation.

In conclusion, we have demonstrated the presence of the TLL transition by “observing’ the variation of the thermodynamic terms of the Thermo-Vogel-Fulcher equation with M via an observer T1, which we define to keep the free volume constant across the values of M, when M varies. We saw that TLL corresponds to the value of T1 that makes the Thermo-Vogel-Fulcher Enthalpy constant as M varies, equal to a characteristic of the melt that we call DHLL. We derived several equations linking the state of the melt, characterized by its value of DHA(M,A), DSA(M,A), to the state of the melt at TLL: DHLL, DSLL=DHLL/TLL, and we saw that the free volume at TLL, ALL,  is constant for all values of M.

           In the next 50 pages of the book's chapter 6 (not expanded in this blog post), it is shown how the presence of TLL influences the rheology of the melt, and is not a “mathematical transition” due to the use of an observer. The reasoning is that since the free volume above and below TLL are different, in the sense that some transition occurs at TLL that modifies the mechanism that generates the free volume, the Newtonian viscosity temperature dependence must be affected when crossing the TLL transition. Consequently, it must be necessary, when performing a regression analysis of Newtonian viscosity-temperature data, for a given M, to distinguish two regions: the one below TLL and the one above TLL, and we should obtain different parameters for, say, the regression of the Vogel-Fulcher equation, Eq. (1) applied to these 2 temperature regions.

          Also, in this same chapter 6, the crucial impact of TLL in determining the melt structural differences below and above TLL is quantified: I quantified the amount of change of free volume at TLL and also the amount of enthalpy change at TLL. I explained, and verified what I had found experimentally: why shear-thinning and the time-temperature shift factors are different below and above TLL and the impact this has on the instability of the melt, leading to “smart processing” capabilities (this is covered in chapter 8 of the Hanser green book pictured above). 

In summary, Boyer was right concerning the existence of TLL. 

            He was also right when he intuitively insisted on its physical importance.  Yet, he was wrong in his view that its cause was linked to a local order explanation, meaning something reminiscent to what initiates crystallization. No, the restructuration of the free volume at TLL  for M < Mc is linked to the coupling between the compensating driving forces of the dual-phase model: [(b/F) ↔ (c,g,t)], a very different reason than a local order explanation. 

The Dual-Phase [b/F ↔(c,g,t)] interpretation of the TLL transition.

We suggest that the TLL manifestation is due to the duality of [b/F ↔(c,g,F)]. We further suggest that the TVF functions, DH(M,T) and DS(M,T), reflect the state of the duality.
The b/F transition controls the nucleation and growth of the b-grains, and their stability, which is a function of the conformation of the conformers participating in the b-grains, itself controlled by the (c,g,t) kinetics. When the thermal energy (kT in the Boltzman’s statistics) evens out the probability to form or melt b-grains, the existence of the dual density fluctuation dissolves in the thermal agitation, releasing the free volume that matched the density difference between the F and the b-conformers.  The F and the b-conformers become locally undistinguishable. Instead of calling this process “the melting of the b-grains”, we prefer to see it as a delocalization of the b-grains, referring to the impossibility to localize the b-grains on the macromolecules. The b-grains melt and reform at different places and with a rate faster than the frequency of observation. One could easily conclude that TLL is the state where the dual-phase physics, modulated by the Grain-Field Statistics, intercepts and fuses with the classical “homogeneous” phase physics, the arena of the Boltzmann’s statistics. Notice that the b-grains continue to form above TLL, yet they are delocalized, hence the presence of the b-grains and the absence of the b-grains is locally indistinguishable. This delocalization results in an increase of the apparent free volume.

At the same time, the increase of free volume from within the macro-coil allows a better inter-penetration between adjacent coils, translating into more bond-sharing of the 2nd kind, or even of a higher kind, in the composition of the b-grains; as a result the entropy increases (the macro-coil radius expands a bit).  The enthalpy continues to increase because the number of conformers, and thus of the formation of new b and F conformers, increases with M: this is just a scaling effect.

 In summary, for T >TLL, the melting of b-grains of the 1st kind, as TLL increases, releases the chain into more reconfiguration options within a macro-coil, not only slightly expanding it, but also creating new (indistinguishable) b-grains, increasing their number as the chain length increases, and favoring their contact sharing with adjacent chains: we assume that this mechanism may be the reason for the increase of entropy for T> TLL for points 7-10. (not shown in this post). We will return to this issue in Chapter 7 of the book.

When T < TLL, for M > Mmc, there is not enough thermal energy “to melt” and delocalize the  existing b-grains and their free volume environment: the rate of formation of the (localized) b-grains wins over their rate of dissociation, yet this happens by a mechanism of growth of the b-grains within the macro-coil, not by increasing their numbers with other chains; this is perhaps why one observes a decrease of the entropy at constant DHLL, by plateau-increments such as 4-5, then 6,7, then 8-10 for the blue dots in Fig. 30 (not shown in this blog post).

Thus my 1st conclusion of this post:
Boyer was right regarding the existence of TLL in the upper melt, although for the wrong reasons.

The 2nd vision of Boyer: the two Tgs in semi-crystalline polymers. Was Boyer also right?  
 
            Not only was I inspired by Boyer to uncover the mystery behind the TLL transition, but I recently (5 years ago) also felt driven to understand the two Tgs of semi-crystalline polymers envisioned by Boyer.

            The question that drove me to undertake this research was even more general: Is the Dual-Phase and Cross-Dual-Phase model applicable to crystallization?

            But first, why do some melts crystallize while others remain amorphous? This should have a simple answer from a dual-phase amorphous model perspective, even if I considered myself primarily focused on the amorphous phase.  If we had two types of amorphous phases, shouldn’t this have an effect on the way the melt crystallizes?

            Crystallization, obviously, is an important research topic which requires a lot of dedication away from the study of the amorphous matter. But crystallization is generated from the amorphous state: I needed to get myself involved!

            I started my own inquiry by studying the influence of entanglement stability on the crystallization process, which led to several lectures and Video Clip Lectures. I published in 2013 a short paper on crystallization from the melt along those lines entitled:
 Crystallization Kinetics for the Split Dual Phase Model of the Amorphous State of Polymers[7]

            The reception of this paper by my peers has not stopped astonishing me: it has become the most read paper of the few I have posted on Research Gate, with more than 2,700 reads in 3 years! This interest pushed me to become more involved in determining whether my model of duality-cross duality could help tackle crystallization.

I can only say that the concept of crystallization emerges naturally from the treatment of the amorphous state and its dual-phase or cross-dual-phase structure. When conformers with the same stable conformation create the equivalent of a b-grain, it becomes the nucleus for crystallization growth. I call it the st-t (stabilized-trans) conformation state, the most stable state achievable. For certain polymers, the classical semi-crystalline polymers, that state is preponderant and easily accessible; for other polymers, it not stable (amorphous polymers). 

Therefore, in my new approach to crystallization, I treat nucleation as a process that allows to pump or release trans conformers from/into the global system Grain-Field statistics: it’s a competing mechanism to b-grain formation: a new duality.

Another idea emerged from this research: to treat crystallization like a cross-duality of the duality [b/F ↔(t, cgf, st-t)]. With that perspective in mind, the crystallization dynamics is treated by analogy with the dual-phase/cross-dual-phase split to explain the formation of entanglements. In the case of the dual-phase-cross-dual-phase compensation, the split allowed the formation of two types of crystals: the micelle type (the M-phase) and the spherulite type (the S-phase), hence two types of amorphous phases, characterized by their Tg(M) and Tg(S), which I associate with Boyer’s Tg(L) and Tg(U).

 In this new approach, crystallization is the result of a competition at two cross-levels: the formation of b-grains within both the S and M phases, and the compensation between the S and M phases. The new model can be quantified by the same equations of the Grain-Field Statistics that drive the interactions between conformers to lead to entanglements.

The idea explored is that the same statistical phenomenon that generates “entanglement” when M increases, - a duality that triggers a cross-duality to minimize the dissipation energy of the global system-, is also at work to produce the 2 amorphous phases, the S and M phases that are themselves dual-phase-coupled with the 2 types of crystalline phases, the S and M crystal-phases!  

The 2nd conclusion of this post is that Boyer was also right regarding the presence of the Tg(L) and the Tg(U) in semi-crystalline polymer melts.

Now, this 2nd aspect of my Boyer’s inspired research might have broader consequences, touching areas other than those I initially considered:

-        The effect of extremely low dilution of a component on the structure of a liquid (isn’t this the current problem challenging the scientific basis of homeopathy?).

-         The Dual-Phase and Cross-Dual-Phase coupling of the electromagnetic field and the gravitation field, and its consequence in understanding the time-space duality and the time-space-inertia cross-duality?
   
The organization of the interactions into M and S cross-dual phases, contemplated for the crystallization from the amorphous phase of polymers, may look like a general idea applicable to all kinds of interactions, and, provided it can correctly be mathematically modeled, I suggest that it is. Simplicity hides itself by way of splitting and “folding” dual-phase solutions of interactions, creating a ladder of entangled systems operating at multiplexed levels; this makes the interactions look more complicated than they actually are!  The cradle for this boundary-less dynamic interlocked network is the electromagnetic field, the alpha-layer: the field without inertia, the medium that interconnects it all after birthing it all. 

 ACKNOWLEDMENT

            I dedicate my findings regarding the QED TLL and the 2 Tgs in semi-crystalline polymers to the memory of Prof. R.F. Boyer (1910-1993), this visionary giant who guided my1st steps in this field.

“ Eh, Ray, despite my crusty car and the bad weather conditions, did we not make it?

            POST-SCRIPTUM.

A new book presenting a dual-phase and cross-dual-phase approach to nucleation and crystallization under dynamic conditions is in the works: “The Dual-Phase and Cross-Dual-Phase Approach to Crystallization in Polymers (writing finished in 2020?).

REFERENCES

1. Raymond F. Boyer (1973), “An apparent double glass transition in semicrystalline polymers”, Journal of Macromolecular Science, Part B, Phys., 8:3-4, 503-537, DOI: 10.1080/00222347308201635

2. Raymond F. Boyer (1963) “The Relation of Transition Temperatures to Chemical Structure in High Polymers”. Rubber Chemistry and Technology: November 1963, Vol. 36, No. 5, pp. 1303-1421. https://doi.org/10.5254/1.3539649

2a. S.J. Stadnicki, J.K. Gillham, and R.F. Boyer, J.Appl. Polym. Sci., 20,1245(1976)

            2b. J.K. Gillham and R.F. Boyer, J.Macromolecular. Sci., Part B, Phys.,13,497 (1977).

            2c J. K. Gillham, Torsional Braid Analysis (TBA) of Polymers, Developments in Polymer Characterisation—3, 10.1007/978-94-009-7346-6_5, (159-227), (1982).

              [3] Jean Pierre Ibar, “ A Theory for the Properties and the Behavior of Polymeric Materials”, June 75.

              [4] J.P. Ibar, (1981) “Non-Newtonian flow behavior of amorphous polymers in the T>T g temperature range: A new analysis of the data according to the “double-shift” procedure”, J.Macromol. Sci., Part B, Physics 19:2, 269 308, DOI: 10.1080/00222348108223943

              [5] This does not mean that crystallization and/or the tendency to crystallize does not affect TLL: it does, but local order is not responsible for TLL in my explanation of it.

            [6.] J.C. MAJESTE (UPPA, PhD thesis, PAU-France).

          [7] J. P. Ibar (2013) Crystallization Kinetics for the Split Dual Phase Model of the Amorphous State of Polymers, Journal of Macromolecular Science, Part B: Physics, 52:7, 950-963, DOI:10.1080/00222348.2012.746571 To link to this article: http://dx.doi.org/10.1080/00222348.2012.746571


Jean Pierre Ibar
October, 2019